More than 3,000 Philips fluoroscopy systems affected by FDA recall
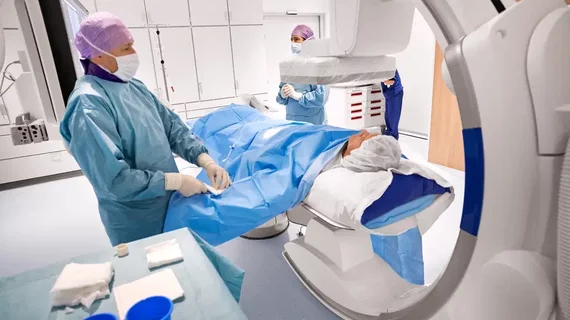
More than 3,000 of Philips’ interventional fluoroscopy systems are the subject of a new recall initiated by the U.S. Food and Drug Administration.
The Class 2 recall pertains to Azurion-brand imaging products, with improperly installed drives potentially impacting the systems’ functionality.
“Solid-state drives (SSDs) that are not specified in the medical device file may have been installed in various PCs,” a notice from the FDA reads. “These SSDs may have different performance characteristics, which may include reduced reliability over time such as loss of imaging functionality or motorized movement, a blank or black display, and/or the inability to store newly acquired images or access previously acquired images.”
On Oct. 6, Philips issued Urgent Medical Device letters to affected customers. The letters informed users that the systems may still be used and included information on workarounds in the event of an issue.
That guidance includes the following:
If motorized movements become unavailable, the stand can be manually repositioned using the handgrips and brake controls located on both sides of the stand.
For ceiling-mounted systems, manual longitudinal, transversal and rotational stand movements remain available. However, transversal stand movements are only available for ceiling-mounted systems with the FlexMove option.
For floor-mounted systems, only rotational stand movements remain available.
The table's longitudinal position can be adjusted manually when the table is not tilted.
Philips intends to inspect the systems and replace the solid-state drives that are not specified in the medical device file, with corrections scheduled to begin in December. Field representatives will be in contact with affected customers to coordinate the repairs.
For more details, click here.
In addition to her background in journalism, Hannah also has patient-facing experience in clinical settings, having spent more than 12 years working as a registered rad tech. She began covering the medical imaging industry for Innovate Healthcare in 2021.